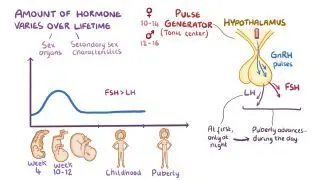
There are two main types of hypogonadism: primary and secondary. Primary hypogonadism is caused by dysfunction of the gonads and can be acquired or congenital. Acquired causes include radiation therapy, chemotherapy, autoimmunity, trauma to the gonads, and certain infections, such as mumps orchitis. On the other hand, congenital causes include genetic disorders affecting gonadal function such as Klinefelter syndrome or Turner syndrome. Regardless of the cause, primary hypogonadism is characterized by a decrease or complete absence of sex hormones, which means there is no negative feedback on the hypothalamic-pituitary-gonadal axis. This leads to an overproduction of gonadotropins (LH and FSH), which is why primary hypogonadism is also known as hypergonadotropic hypogonadism.
On the other hand, secondary hypogonadism, also known as hypogonadotropic hypogonadism, is characterized by low levels of LH and FSH. Secondary hypogonadism occurs due to hypothalamus or pituitary gland dysfunction causing impaired GnRH or gonadotropin secretion. Acquired causes of secondary hypogonadism include tumors of the pituitary gland and hypothalamus; pituitary apoplexy (i.e., sudden bleeding or impaired blood supply to the pituitary gland); and trauma to the base of the skull. Congenital causes include panhypopituitarism, which involves a decrease in all pituitary hormones, as well as Kallmann syndrome, a genetic condition marked by isolated GnRH deficiency and anosmia or hyposmia (i.e., complete or partial loss of the sense of smell). Finally, general causes of secondary hypogonadism include conditions that affect hormonal release by the hypothalamus and pituitary gland, such as chronic illness (e.g., cystic fibrosis or celiac disease), malnutrition or obesity, excessive exercise, and stress.